Peptide Mass Spectrometry | Molecular Weight and Identity Checks
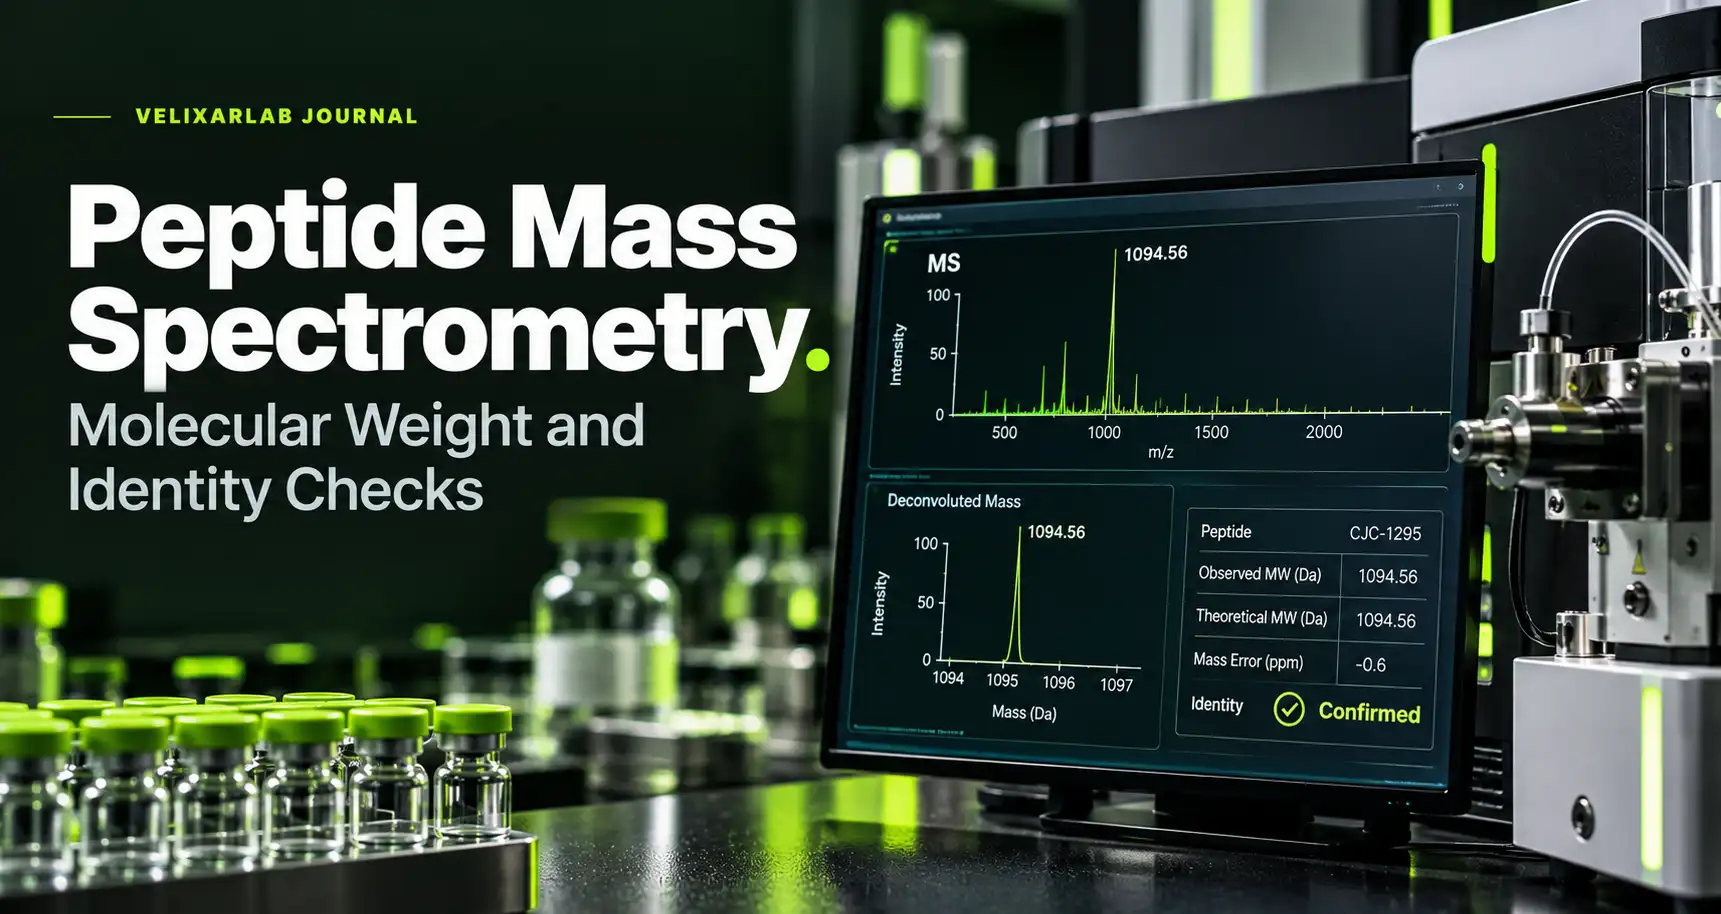
- Mass spectrometry measures mass-to-charge information. It detects ions, not a molecule’s name printed on a product label.
- Peptides may carry several charges. An electrospray spectrum can show several m/z peaks belonging to one molecular species.
- Deconvolution helps recover molecular mass. Theoretical and observed masses can be compared after accounting for charges and relevant adducts.
- An intact-mass match supports identity, but has limits. Different sequences can share a mass, and stereoisomers may not be distinguished by mass alone.
- LC-MS adds separation before mass detection. It can improve assessment of components, but co-elution and unequal ionisation remain considerations.
- Purity and quantity are separate measurements. A mass signal does not automatically establish HPLC area purity or net peptide content per vial.
- A useful COA must trace to the actual lot. Verify sample identifiers, instrument method, expected and observed values and issuing laboratory.
A chromatogram may display one dominant peak, while a mass spectrum contains several signals. Does that mean the sample contains several different peptides? Not necessarily. Peptide mass spectrometry identity work begins by distinguishing the neutral molecular mass from the mass-to-charge values of the ions actually measured.
For laboratories handling analytical research materials in India, peptide mass spectrometry identity provides one important line of evidence about the molecular species in a submitted sample. It should be interpreted alongside chemical specifications, suitable chromatography and lot traceability rather than used as a universal certificate of quality.
What Does a Mass Spectrometer Measure?
For peptide mass spectrometry identity reporting, instrument mass accuracy should not be treated as the sole measure of analytical certainty. An accurate measurement can still support the wrong assignment if the sample chemistry, charge state or candidate list is incomplete.
A mass spectrometer ionises compounds and separates or detects the resulting ions according to their mass-to-charge ratio, written *m/z*. The output is a spectrum, usually showing signal intensity against m/z. The instrument does not read the product name or directly weigh the entire vial.
For peptide mass spectrometry identity, a laboratory starts with a theoretical mass calculated from a specified amino-acid sequence, including applicable chemical modifications and termini. The measured ion pattern is then examined to determine whether it is consistent with that proposed species.
Average Mass Versus Monoisotopic Mass
Different mass calculations may be used. *Monoisotopic mass* is based on the most abundant specific isotopes of each element, whereas *average molecular mass* incorporates natural isotope distributions. A reference value and an instrument report must be compared using compatible definitions.
If a theoretical mass and an observed mass differ, the reviewer should first check whether they refer to the same species and mass convention. Salt form, adducts, chemical modifications, oxidation and calibration can also affect interpretation.
Why One Peptide Can Produce Several m/z Peaks
A peptide mass spectrometry identity report may show an isotope envelope around each charge-state signal. The spacing of isotope peaks can help infer charge, while the relative pattern may support—but not uniquely guarantee—the chemical assignment.
Electrospray ionisation commonly produces multiply charged peptide ions. A single peptide may appear as a family of signals corresponding to ions with one, two, three or more positive charges, depending on molecular features and experimental conditions.
A simple positive-ion relation is m/z ≈ (M + z × H) / z, where *M* is neutral molecular mass, *z* is charge number, and *H* is the mass of a proton. This conceptual formula is useful, but actual interpretation may also require accounting for adducts, isotope patterns and instrumental accuracy.
In peptide mass spectrometry identity, the presence of three apparent peaks does not automatically mean three impurities. Analysts may use charge-state series and isotopic spacing to connect them to one molecule.
Fictional Charge-State Example
Imagine a fictional peptide with a neutral mass near 3,000 Da. Depending on its charge state, protonated ions could appear around m/z 1,501 for +2 and m/z 1,001 for +3. These round values illustrate the concept; they are not actual data from a VelixarLab certificate. A qualified analyst would use precise mass constants and the instrument’s measured values.
The ability to identify a related charge-state family is central to reliable peptide mass spectrometry identity work.
LC-MS Versus Mass Spectrometry Alone
Peptide mass spectrometry identity assessment benefits from comparing chromatographic retention with mass information. Nevertheless, a broad LC peak may contain co-eluting species, and a mass detector may respond to chemically different analytes with unequal efficiency.
*LC-MS* means that liquid chromatography is coupled to mass spectrometry. The liquid-chromatography stage separates compounds over time; the detector then records m/z signals for components as they elute. This can help distinguish multiple chemical species in complex samples, provided the chromatographic and mass-spectrometric methods are suitable.
A well-designed peptide mass spectrometry identity workflow can use an extracted-ion chromatogram, a total-ion chromatogram, an intact-mass spectrum and, where appropriate, tandem MS. Each output answers a slightly different question.
| Technique / output | Useful question | Key limitation |
|---|---|---|
| LC retention time | When does the component elute? | Not unique identity by itself |
| Accurate intact mass | Is observed mass consistent with target? | Isomers may have identical mass |
| Isotope pattern | Is elemental composition plausible? | Interpretation needs adequate resolving power |
| MS/MS fragments | Do product ions support the sequence? | Coverage and ambiguity depend on the experiment |
| HPLC UV area % | What fraction of included detector area is assigned to a peak? | Not the same as mass fraction or sequence identity |
| Calibrated assay | How much target is present? | Depends on standards, response and validation |
See the for introductory instrument concepts. This is background methodology, not independent authentication of a particular supplier’s material.
How a Laboratory Checks a Theoretical Peptide Mass
In peptide mass spectrometry identity work, calculate the theoretical mass from the complete sequence and all stated terminal modifications, then compare it with the correct ion type. A product’s counterion or hydrate may affect the mass of sold powder without being the neutral peptide molecular mass used for MS identity.
First identify the exact sequence and any chemical modifications. N-terminal acetylation, C-terminal amidation, oxidation, disulfide bonds and other changes can affect predicted mass. A peptide’s salt or hydration state may matter separately to gross powder weight.
Next, determine whether the report’s observed signals align with the expected charge states and mass type. Ensure the instrument method was suitable for the intended identity question. Peptide mass spectrometry identity is strongest when the report documents the expected and observed values, relevant tolerance and how the main species was assigned.
A historic illustrates that charge-state analysis and mass accuracy can be used to characterize biologically active peptides. The instrument’s mass accuracy must be interpreted in the context of that method, not assumed from a general manufacturer’s specification.
What If the Observed Mass Does Not Match?
Possible explanations include a different sequence, a terminal-group discrepancy, chemical modification, adduct formation, degradation, incorrect charge assignment or a reference/calibration issue. Do not automatically attribute the difference to fraud; preserve the report and ask the issuing laboratory to explain the assignment with data.
For peptide mass spectrometry identity, a mismatch is an analytical question that needs resolution before the material is treated as fully identified for a demanding research protocol.
Why Intact Mass Does Not Prove Complete Sequence Identity
For a higher-confidence peptide mass spectrometry identity conclusion, an institution may need a second analytical dimension such as fragmentation coverage, peptide mapping or a reference sample. The chosen evidence should match the scientific risk and not simply the equipment available.
Two different sequences can have the same elemental composition and therefore the same exact mass. Leucine and isoleucine, for example, are isomeric amino acids with the same elemental formula. A peptide containing one rather than the other may not be distinguishable using intact mass alone.
That limitation makes peptide mass spectrometry identity an exercise in combining evidence. Tandem MS, chemical or enzymatic peptide mapping, chromatographic reference comparisons and other suitable techniques may be needed for some structures. Even a strong MS/MS assignment has limitations when fragmentation is incomplete or two candidates yield similar ions.
The specifically frames peptide mapping as an identification technique in appropriate contexts.
How to Read an Identity Section on a Research COA
A peptide mass spectrometry identity note should explicitly state whether the result is “consistent with expected mass” or “identity confirmed by a validated method,” as those claims require different support. Laboratory auditors should be able to find the related acceptance criterion and raw-data reference.
| COA field | What a reviewer needs |
|---|---|
| Material specification | Sequence, declared modifications and chemical form |
| Sample or lot ID | Link to the supplied vial or bulk lot |
| Instrument method | LC-MS, HRMS, MS/MS or another stated technique |
| Theoretical value | Appropriate neutral / ion mass and mass definition |
| Observed result | Recorded values and accepted measurement tolerance |
| Conclusion | Precisely what the analytical evidence supports |
| Issuing laboratory | Traceable report number, date and authentication route |
In peptide mass spectrometry identity documentation, terms such as “mass confirmed” are too vague when the research requires sequence-level confidence. Ask whether the lab measured intact mass, fragments, a reference-standard comparison or a validated identity procedure.
Do not use a product listing’s “99% purity” claim as a stand-in for the expected and observed molecular masses. Purity, identity and vial quantity should have distinct supporting results.
Examples From VelixarLab’s Research Catalogue
VelixarLab India provides research-product descriptions for and . These illustrate why identity methods need to fit the target. A 16-residue mitochondrial-derived peptide and a copper-associated tripeptide involve different analytical questions; a copper complex may need metal-speciation evidence in addition to peptide identification.
A review of peptide mass spectrometry identity should be linked to the actual proposed material and batch, not a generic statement about all research products. Ask for available documentation through before deciding if the sample meets your institution’s requirements.
Common Misreadings of Mass Spectrometry
Peptide mass spectrometry identity methods are powerful when they are transparent about uncertainty. A missing or unusual signal warrants method-aware investigation, not automatic reclassification of a compound or confident speculation about its impurity profile.
- Calling every charged ion peak a separate impurity.
- Comparing an average theoretical mass with a monoisotopic observed mass without explanation.
- Assuming intact-mass agreement proves amino-acid order or stereochemistry.
- Dividing uncalibrated MS signal areas to infer molar ratios or mg per vial.
- Treating sample preparation artifacts or adducts as verified parent compound species.
- Using a genuine report that belongs to a different batch.
The aim of peptide mass spectrometry identity review is not to demand the maximum number of tests, but to match each analytical claim to an appropriate method.
Frequently Asked Questions
What Does m/z Mean in Peptide Mass Spectrometry?
It means mass-to-charge ratio for an ion. A multi-charged peptide can appear at several m/z values.
Is Molecular Weight the Same as m/z?
Not generally. Molecular mass describes a neutral chemical species; m/z also depends on charge and added or removed ions.
Does LC-MS Prove Peptide Purity?
It can help characterize mixtures and impurities under a suitable method, but an identity result is not automatically a validated quantitative purity or content assay.
Can Mass Spectrometry Distinguish Leucine and Isoleucine?
Not by intact mass alone, because the two residues have the same elemental composition. Additional methods may be needed.
Is One Matching Mass Peak Enough for a COA?
It may provide useful evidence, but adequacy depends on the required identity confidence, method and chemical alternatives.
What Should an Indian Laboratory Request?
A sample-linked report naming the method, exact species, expected and observed values, acceptance criteria where applicable and the responsible laboratory.
Conclusion: Read the Ions, Not Just the Headline
Peptide mass spectrometry identity is powerful because it connects chemical specifications to experimentally measured ion data. But correct interpretation requires charge states, mass conventions, possible adducts, sequence ambiguities and method limits to be understood. The best research documentation integrates identity with chromatographic and quantitative evidence while preserving lot traceability.
For a particular peptide material, start with and for the exact lot under consideration.
*Research-use notice: This technical article covers analytical chemistry, not personal administration, therapeutic effect or approval of experimental compounds.*
References — scientific and technical sources
- D’Agostino et al. (1997), bioactive peptides by LC-high resolution electrospray MS
- USP <1055>, peptide mapping for confirmation of identity
- ICH Q2(R2), analytical procedures for identification, purity and assay
- Agilent, fundamentals of liquid chromatography–mass spectrometry